
CRISPR & Prime Editing SMARTSeq
Price range: 40,00€ through 190,00€
Select the number of each type of sample you want to sequence.
Description
All-in-one CRISPR & Prime Editing analysis
CRISPR & Prime Editing SMARTSeq combines Long Read sequencing with our dedicated CRISPR / Prime Editing bioinformatics pipeline: Amplicon Long Read SMARTSeq for a homogeneous amplicon from an isolated clone, or DNA Pool Long Read SMARTSeq for a pool of amplicons. Full length amplicons are sequenced: indels, knock-ins and multi-site edits are seen together on the same molecule.
Which edits do we detect?
- Knock-out – insertions and deletions at the cut site, with size and position of each event (NHEJ indel rate).
- Knock-in / HDR – exact or near-exact integration of your template versus WT and indel reads (HDR rate).
- Prime Editing – all expected substitutions checked together on the same read: complete vs partial editing, compared with a WT control.
- Multi-guide designs & edited clones – editing rate per gRNA (each cut site analysed in its own window) and allele-by-allele phasing of clones.
- Motifs on both strands – sequence diversity around junction motifs on the top and bottom strands.
Which sequencing depth should I choose?
- Up to 1,000 reads – isolated clone – one amplicon from an isolated clone, to confirm the presence and nature of the edit on each allele.
- Up to 5,000 reads – amplicon pool – pooled amplicons from an edited cell population, single knock-out or knock-in.
- Up to 10,000 reads – amplicon pool – edited cell pools, several guides on the same amplicon, mixed populations.
- Up to 15,000 reads – amplicon pool – low-frequency events, such as prime editing in DNA pools.
What you receive in your Cloud Space
- HTML report, generated only when all quality checks pass
- Excel workbook: read counts, percentages and event signatures
- FASTQ files sorted by read category (WT, edited, indel, ambiguous…)
- BAM + BAI files for visualisation, with a step-by-step viewing guide
- QC validation workbook
- Methods and parameters ready for your publication
Sample requirements
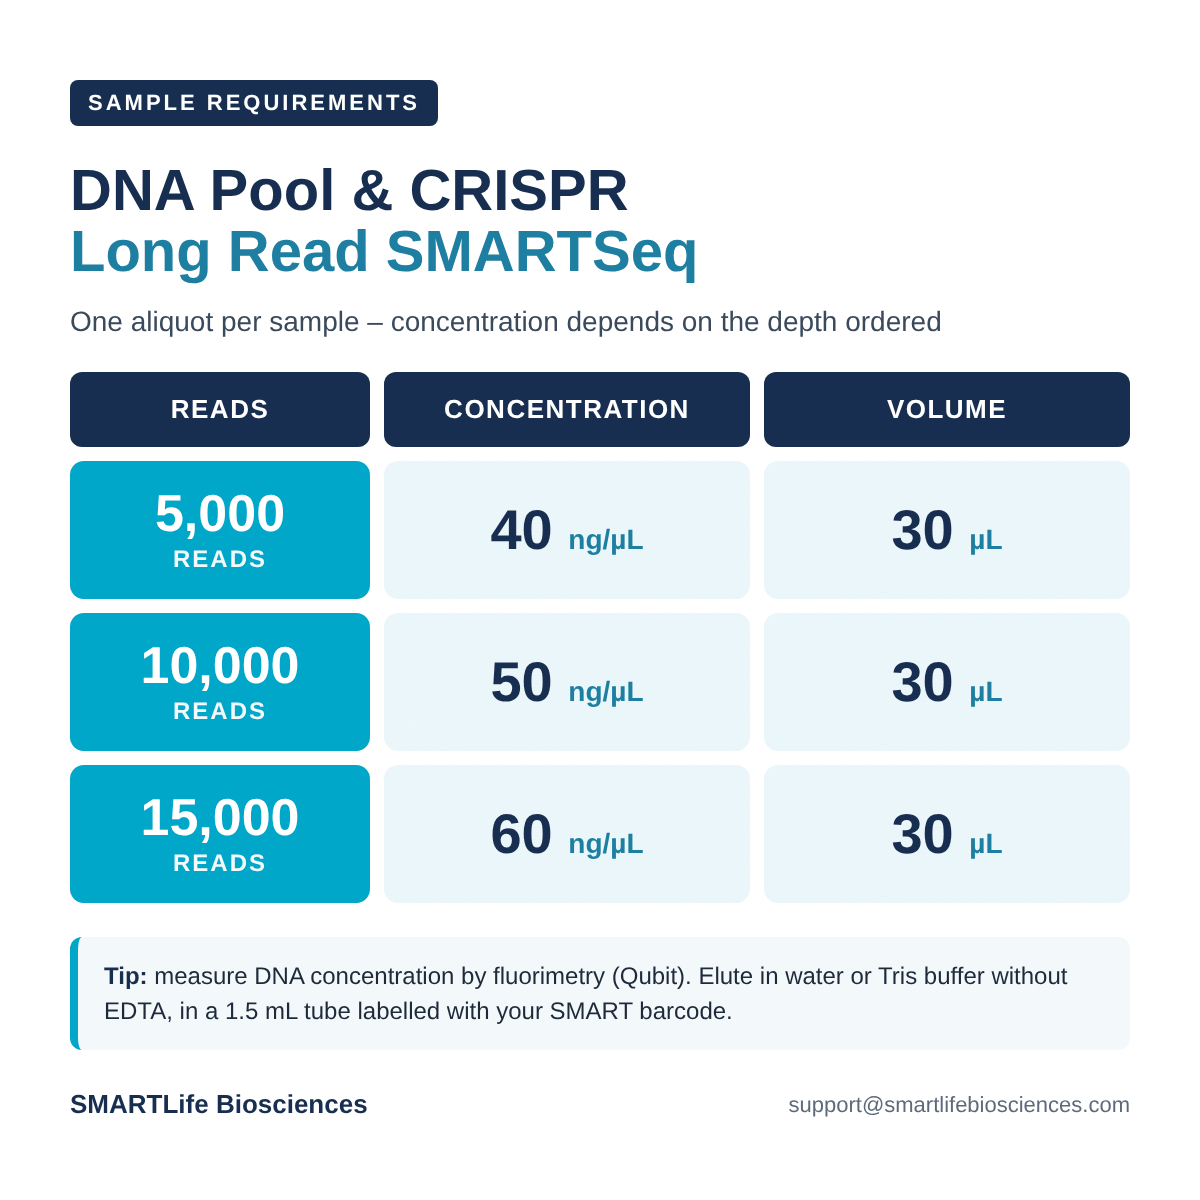
| Reads | Sample | Concentration | Volume |
|---|---|---|---|
| Up to 1,000 | Isolated clone – 1 amplicon | 30 ng/µL | 30 µL |
| Up to 5,000 | Amplicon pool | 40 ng/µL | 30 µL |
| Up to 10,000 | Amplicon pool | 50 ng/µL | 30 µL |
| Up to 15,000 | Amplicon pool | 60 ng/µL | 30 µL |
Design recommendations
- Position your cut site at least 100 bp from each end of the amplicon. Our routine sequencing uses a Rapid Barcoding (transposase-based) library preparation, which reduces coverage at the very ends of the amplicons. Editing is analysed in a window of ±30 bp around the cut site: keeping the cut site 100 bp away from the ends ensures that this whole window is covered by enough full-quality reads.
- Several gRNAs on the same amplicon: each cut site is analysed in its own ±30 bp window. Apply the same 100 bp margin to the first and last gRNA.
- Isolated clone: send a single amplicon from one clone. The reads are clustered by allele, so heterozygous or compound edits are resolved allele by allele.
- Amplicon pool: amplicons from an edited cell population or from several samples pooled in one tube. Whenever possible, normalize each amplicon before pooling so that all targets receive enough reads.
- Cut site closer than 100 bp to an amplicon end? Please mention it in the remark section of your order and contact our support team before sending your samples: support@smartlifebiosciences.com. We will review your design with you and advise on the best option.
Recommended purification methods
- Magnetic bead-based purification: AMPure XP beads (or equivalent SPRI magnetic beads such as SPRIselect), NEB Monarch Mag PCR & DNA Cleanup Kit
- Column-based kits: QIAquick PCR Purification Kit (Qiagen), Monarch PCR & DNA Cleanup Kit (NEB), NucleoSpin Gel and PCR Clean-up (Macherey-Nagel)
To preserve DNA integrity
- Do not vortex DNA samples
- Mix gently by slow pipetting or tube inversion
- Avoid vigorous shaking
- Avoid repeated freeze-thaw cycles
- Store DNA in nuclease-free water or 10 mM Tris-HCl pH 8.0
- Minimize exposure to high temperatures
Contaminants to avoid
- Residual ethanol, phenol or isopropanol
- EDTA (high concentrations)
- Guanidinium salts or excess salts
- SDS, CTAB, Triton, Tween or other detergents
- Protein or RNA contamination
- Primers, primer dimers or residual dNTPs
Quality control recommendations
- Quantification: Qubit dsDNA HS or BR Assay – avoid relying solely on NanoDrop measurements
- Whenever possible, verify the absence of primer dimers by gel electrophoresis, TapeStation or Bioanalyzer
- Amplicon pools: whenever possible, normalize individual amplicons before pooling
What to send
- Your sample, up to 25 kb – one amplicon from an isolated clone, or an amplicon pool (see sample requirements above)
- To upload in FASTA format when placing your online order:
- WT reference sequence
- Expected edited sequence(s): knock-in, prime editing signature…
- gRNA / pegRNA sequences
- In the remark section of your order: confirm the nuclease used (Cas9 or any other Cas enzyme)
- An unedited control sample is recommended to measure background
A specific editing design (multiplexed guides, large knock-ins, rare events in pools)? Contact our team: support@smartlifebiosciences.com